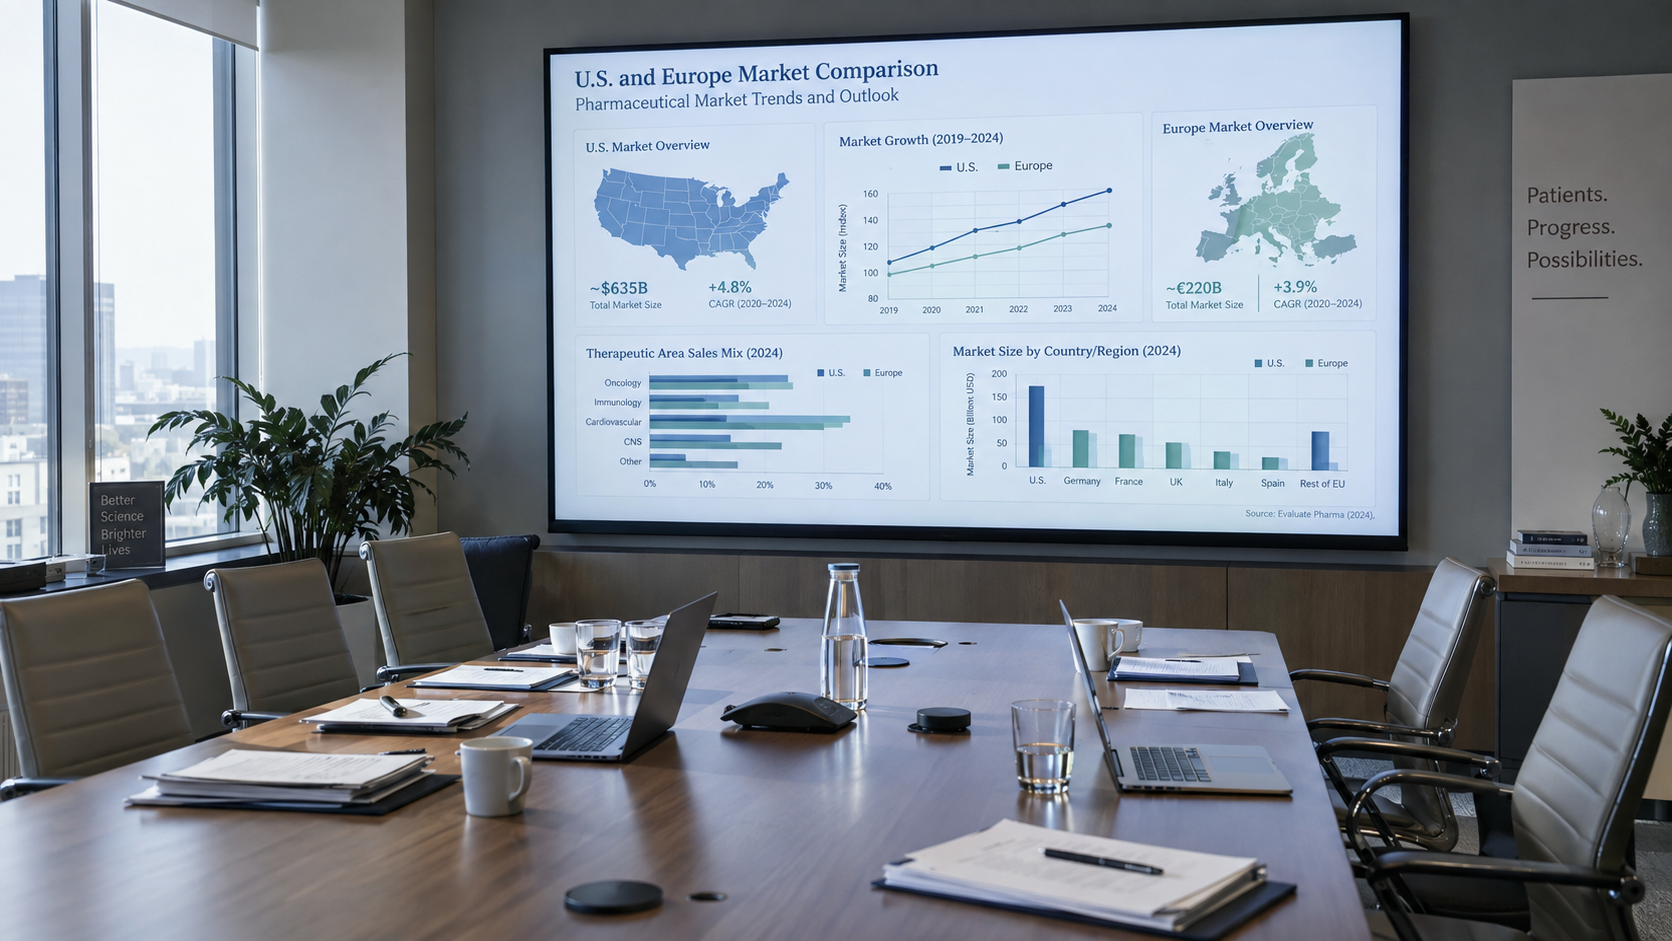
Most-Favored-Nation Pricing Is Becoming a Global Management Problem
The expansion of the GENEROUS Medicaid model and newly disclosed terms in Pfizer’s U.S. pricing agreement show how decisions about pharmaceutical pricing outside the United States can increasingly affect U.S. reimbursement and company economics. Global pricing decisions may need to be managed with a much wider lens.
For pharmaceutical companies, the latest U.S. drug-pricing developments are getting harder to look at one at a time.
President Trump announced that all 50 states, Washington, D.C. and Puerto Rico intend to participate in the GENEROUS Medicaid model. The program is designed to give Medicaid access to net prices for certain medicines that are comparable to prices paid in selected foreign countries.
At the same time, newly released contract language involving Pfizer reportedly shows that the company agreed to share with the Department of Health and Human Services a portion of additional net revenue generated when prices for certain medicines increase outside the United States.
Those two developments matter together because they show how pricing decisions made in Europe and other markets can affect U.S. reimbursement and U.S. government agreements.
GENEROUS Brings International Pricing Into Medicaid
The immediate importance of GENEROUS is its reach. Every state has now expressed interest in participating in a model that uses international pricing as part of the way Medicaid rebates are calculated for participating medicines.
That does not mean every drug suddenly gets an international reference price. Manufacturers still have to participate, products will differ, and many of the details matter. But there is enough here for management teams to pay attention.
A price negotiated in Europe or another reference market may affect the economics of the same product in the United States. That makes it harder to treat global pricing as a collection of separate country-by-country decisions.
The Pfizer Agreement Adds Another Layer
The Pfizer disclosure makes the issue more interesting.
According to the reported contract language, Pfizer must share with HHS part of the additional revenue generated when prices for certain medicines rise outside the United States. The public version does not reveal the percentage, the covered medicines or several other important terms.
Even with those gaps, the structure is worth watching.
A higher price overseas could improve revenue in that market while also creating a separate financial obligation in the United States. That is different from a traditional reference-pricing model, where the main concern is how a foreign price may affect a U.S. benchmark.
Here, a commercial decision in one country may create an economic consequence somewhere else.
This Is a Management Issue
That is why this is becoming more than a pricing issue.
The practical management question is whether the company understands which decisions made outside the United States can affect its U.S. economics.
That requires market access, finance, legal, government affairs and commercial teams to work from the same picture. Each group may be making a reasonable decision on its own, but problems can arise when those decisions are made without understanding what they mean for another market.
A locally successful pricing decision can still create an unexpected consequence elsewhere.
What Management Should Be Looking At
Companies should start by mapping their major products against the pricing arrangements that could matter in the United States. Management should understand which international prices could affect GENEROUS or other U.S. pricing arrangements, which products are covered by company-specific agreements, and what happens when the price of a medicine changes in another market.
There should also be a clear process for bringing pricing, legal, finance, government affairs and commercial leadership together before a significant pricing decision is made.
The goal is not to predict every future policy change. It is to make sure a decision in one market does not create an expensive surprise in another.
The Bigger Picture
There is still a lot we do not know. Many of the pharmaceutical agreements being negotiated with the administration remain confidential, and some of the most important commercial terms have not been made public.
But companies already have enough information to see the broader issue.
International pricing is becoming more closely tied to U.S. reimbursement. GENEROUS does that through Medicaid rebates linked to prices in other countries. The Pfizer agreement reportedly adds another connection by tying increases in overseas revenue to payments back to HHS.
For executives, the question is becoming less about what price should be negotiated in a particular country and more about what that decision does to the economics of the product across markets.
That is the issue management teams should be thinking about now.
About Lanton Strategies International & Lanton, Lanton and Sosa Law, PLLC
Lanton Strategies International works with executives and organizations on policy, market access, government affairs and strategic issues affecting investment, commercialization and growth across the United States and Europe. We help leadership teams understand what policy developments mean for their business and determine practical next steps.
For legal and regulatory matters, Lanton, Lanton & Sosa Law PLLC advises companies on issues affecting healthcare, life sciences, technology, commercialization and business strategy.
This article is provided for informational and educational purposes only and should not be relied upon as legal, investment or other professional advice. Reading it does not create an attorney-client relationship.
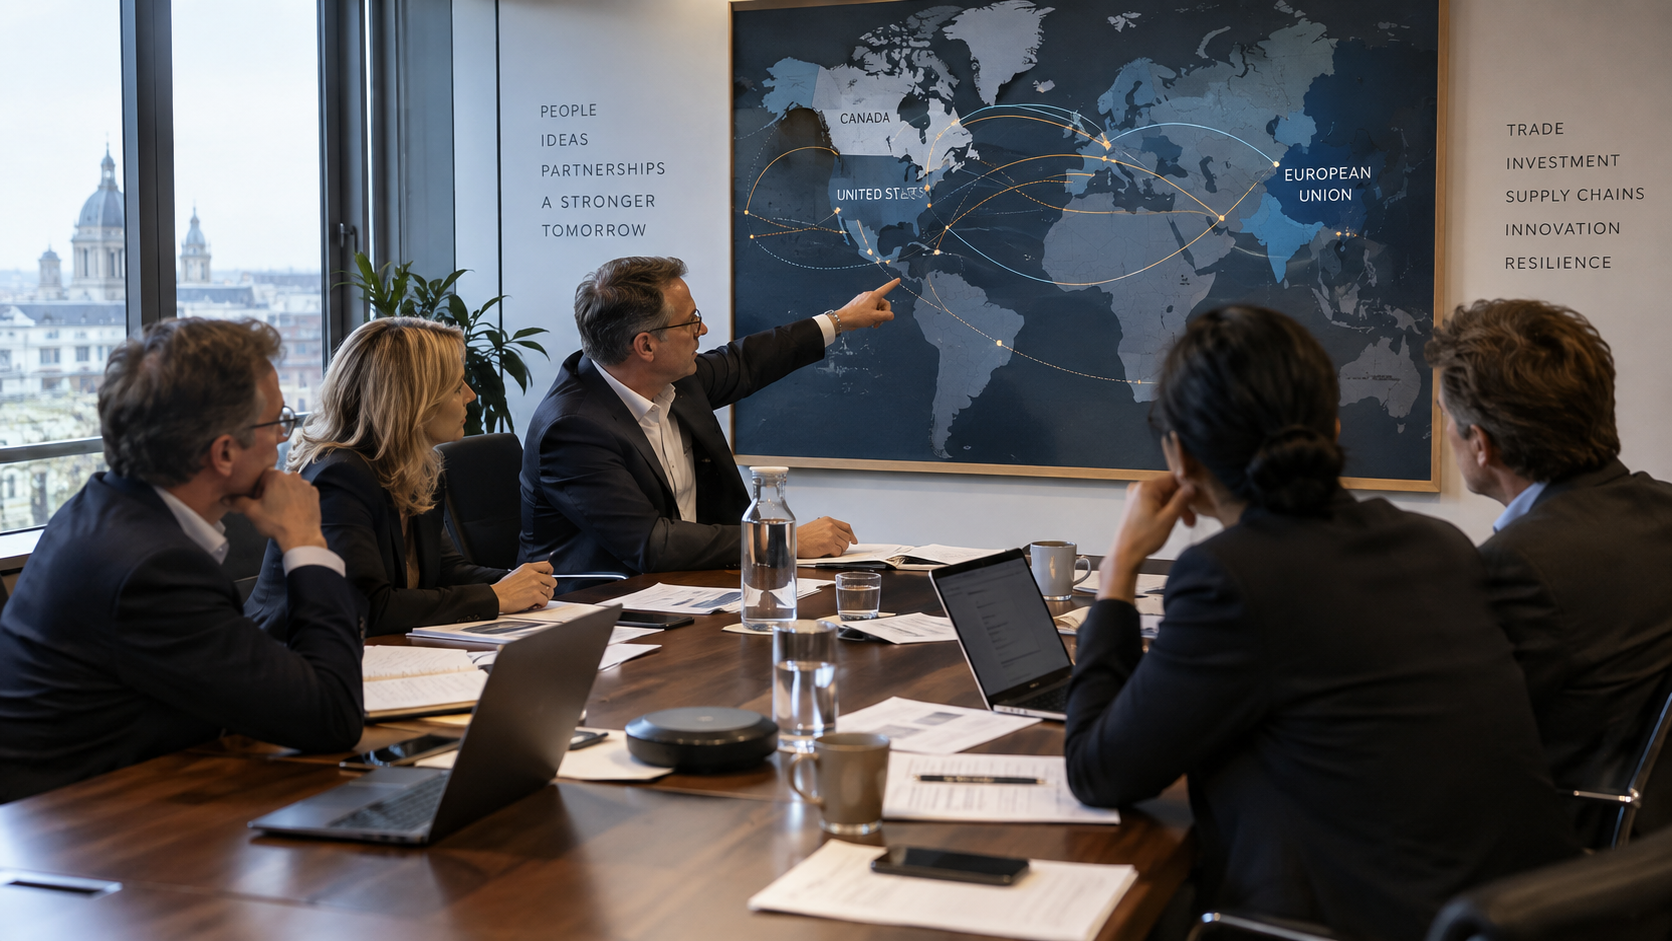
Trump’s New EU Tariff Threat Creates Another Planning Problem for CEOs
President Trump’s latest tariff threat toward Europe adds another layer of uncertainty for companies making decisions about manufacturing, supply chains and cross-border investment. The right response is not panic. It is understanding exposure, testing assumptions and preserving flexibility before uncertainty becomes an expensive business problem.
European executives making decisions about U.S. manufacturing, sourcing and capital deployment have another variable to consider.
On September 17, 2026, President Donald Trump threatened heavier tariffs on Europe if the European Union moves ahead with a proposal to create a closer relationship with Canada. Canadian Prime Minister Mark Carney has called for deeper cooperation with Europe in areas including trade, technology, energy and critical minerals. Reuters Financial Times
The proposed EU–Canada relationship is not fully defined, and there is no new tariff schedule to model today. Companies should not redesign their businesses around every political statement.
They also cannot ignore the possibility that trade conditions between the United States, Europe and Canada could become less predictable. That uncertainty is the real issue.
For a European company considering a U.S. facility, a major sourcing decision or a new market entry, the question is simple:
Which assumptions become expensive if they are wrong?
That is where management should focus. A company should understand how different tariff scenarios could affect the economics of a new investment, where its supply chain is most exposed, which decisions can wait and where flexibility should be preserved. The best response is not panic. It is scenario planning.
The Canada–EU relationship matters even before the details are settled
The EU’s proposal is broader than a conventional trade discussion. The emerging relationship could involve cooperation in technology, healthcare, energy, security, critical minerals, digital trade, research and other strategic areas.
For companies, that may create opportunities as well as risks. Closer Canada–EU cooperation could lead to new partnerships, sourcing options and investment relationships, particularly in healthcare, technology, advanced manufacturing and critical materials.
At the same time, a stronger Canada–EU relationship could produce new questions for businesses that operate across all three markets. Which rules will apply? Will supply chains become more diversified or more complicated? Could political retaliation affect investment decisions, customs treatment or market access?
Those questions require companies to understand the actual structure of their operations and the assumptions behind major commitments.
The business consequence matters more than the political prediction
CEOs do not need to predict whether Canada ultimately receives a new relationship with the EU or whether Washington follows through on additional tariffs.
They need to know what each plausible outcome could mean for the business.
That means identifying exposure across manufacturing, sourcing, logistics, contracts, data, financing, regulatory approvals and customer commitments.
It also means separating decisions that require action now from decisions that should remain flexible until more is known. Political signals can change quickly. Factories, contracts, supply chains and capital commitments cannot.
What executives should review now
Leadership teams should begin by mapping which products, services and investments depend on the United States, Europe or Canada. They should identify where goods cross borders, where important inputs originate and which contracts assume stable tariff or regulatory conditions.
They should also test whether a proposed investment still makes sense under several scenarios. A project that works under current assumptions may look different if tariffs increase, customs treatment changes or a company loses access to a preferred supplier.
The exercise does not require predicting the future. It just simply requires understanding the cost of being incorrect.
Companies should also review whether their contracts address tariff changes, regulatory shifts, supply interruptions, delays, force majeure, price adjustments and termination rights. These provisions may become more important as trade policy becomes less predictable.
A planning problem, not a reason to freeze
Uncertainty can cause companies to delay decisions unnecessarily. That can be just as costly as moving too quickly.
The goal is not to stop investing, entering markets or building relationships. The goal is to understand which decisions are durable, which can be staged and which should preserve room to adapt.
A company may decide to proceed with a facility but phase the investment. It may retain more than one supplier. It may structure a contract with clearer adjustment mechanisms. It may test a market before making a larger commitment. Those are practical and measured responses to uncertainty.
The lesson for CEOs
The current dispute over Canada, the European Union and possible U.S. tariffs is part of a larger change in the business environment. Executives are planning around movement, not around a stable set of trade and regulatory assumptions.
The practical lesson is straightforward: separate the headline from the business consequence.
Understand where the exposure sits. Test the assumptions behind major decisions. Review the contracts that support those decisions. Preserve enough flexibility to move if the policy environment changes.
Lanton Strategies International works with executives and organizations on policy, market access, government affairs and strategic issues affecting investment, commercialization and growth across the United States and Europe. We help leadership teams understand what policy developments mean for their business and determine practical next steps.
For legal and regulatory matters, Lanton, Lanton & Sosa Law PLLC advises companies on issues affecting healthcare, life sciences, technology, commercialization and business strategy.
This article is provided for informational and educational purposes only and should not be relied upon as legal, investment or other professional advice. Reading it does not create an attorney-client relationship.
AI + Healthcare, Part 2: What Happens When the Product Keeps Changing After Approval?
FDA approval is not the end of the regulatory or commercial journey for AI-enabled healthcare products. This article examines what happens when AI changes a medical device, diagnostic tool or clinical software platform after approval—and how companies should manage regulatory review, reimbursement, accountability and investor expectations.
The harder regulatory question may begin after FDA clearance or approval.
A medical device, diagnostic tool or clinical software platform may reach the market based on one version of its technology. The company then uses AI to improve performance, identify new patterns, reduce false positives or expand the product’s usefulness.
That creates a new practical question for leadership: when is an update still part of the approved product, and when has the company created something that needs another regulatory review?
A conventional medical product usually stays recognizable after approval. Its manufacturer may improve the design, but providers, payers and regulators can still identify the product they evaluated and compare the changes against the earlier version.
AI-enabled software can change things more quietly. A new training set, algorithm or software release may alter the product’s recommendations without changing its outward appearance. The company’s process for managing those changes therefore should become part of its regulatory strategy.
For example, an update that fixes a technical problem may raise different questions from one that changes the product’s intended use, clinical claims, risk profile or method of operation. A diagnostic tool that becomes capable of identifying a different condition, guiding a new treatment decision or serving a broader patient population may require more than ordinary maintenance.
Executives should establish those boundaries before the product reaches the market. Waiting until an engineering team has completed a major update can leave the company trying to answer regulatory questions after the commercial decision has already been made.
Approval does not end the regulatory work
AI development often follows a continuous cycle. Teams release improvements in response to new data, customer feedback and performance testing. FDA review, however, is based on defined submissions and defined evidence.
A company needs a practical plan for what happens next. It should know which changes can be handled through an established change-control process, which require additional validation and which may require a new submission. It should also be able to show how the updated product performs across the populations and care settings where it will be used.
This does not require companies to abandon rapid development. It requires technical, clinical and regulatory teams to make decisions together before an update is released.
Payment and adoption create a second gate
FDA authorization allows a product to be marketed under the applicable rules. It does not guarantee coverage, payment or adoption.
An AI update may improve accuracy or expand the number of patients who can benefit. A payer may still ask whether the change improves outcomes, lowers total cost or simply produces a more sophisticated recommendation.
Hospitals may ask whether the update changes workflow, documentation, staffing or liability. Health plans may ask whether the revised product fits within an existing coverage category or requires new evidence.
A company that treats reimbursement as a post-approval issue may discover that its commercial plan has not kept pace with its regulatory progress.
Responsibility follows the update
Leadership also needs a clear answer to a basic question: who is responsible when the product changes?
That answer depends on how the company tests and validates each release, notifies providers and customers, maintains version records and manages human review.
It also depends on cybersecurity, data governance and the contracts that allocate responsibility among the developer, provider and health system.
If the system produces a different result after receiving new data, the company should be able to explain what changed, why it changed and how the revised performance was evaluated.
That is a business requirement as much as a legal one.
The investment question
Investors may see continuous AI improvement as a competitive advantage. They may also ask whether the company has the systems needed to manage continuing FDA, reimbursement, quality and liability obligations.
A company that can show disciplined product governance may be better positioned than one that treats every update as a software release with no regulatory consequences. The quality of that governance can affect launch timing, contracting, payer discussions and investor confidence.
The central issue is not whether AI should continue improving after approval. It should.
The issue is whether the company has built a process that allows improvement without losing track of authorization, evidence, payment and accountability.
FDA approval marks an important milestone. For an AI-enabled product, it may also mark the beginning of a more demanding operating phase.
The companies most likely to succeed will connect engineering, regulatory, clinical, reimbursement and legal decisions before an update reaches the market.
That is the real test for healthcare AI should not be simply whether the product can learn, but whether the company can govern what it learns.
This is the second article in a series examining artificial intelligence through healthcare, regulation, reimbursement, capital and the institutions that determine whether innovation reaches patients.
Lanton Strategies International advises healthcare and life-sciences companies on U.S. market entry, policy, reimbursement, regulatory strategy and commercial risk. Lanton, Lanton & Sosa Law PLLC provides legal counsel to regulated healthcare organizations, associations and businesses.
This article is provided for general educational and informational purposes only. It is not legal advice and does not create an attorney-client relationship. Readers should consult qualified counsel about their specific circumstances.